Study Reveals Best Segmentation Methods for Spatial Transcriptomics Across Diverse Tissues
October 23, 2026


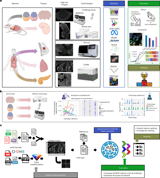
A unified, scalable benchmarking framework evaluated 11 segmentation methods for spatial transcriptomics across 15 datasets from five technologies, covering 12 tissues in humans and mice to see how segmentation influences downstream biology.
The study uses a Nature Methods benchmark to compare methods and understand how segmentation changes conclusions about biology across diverse platforms and tissues.
Trade-offs emerged: SAM and SAM2 offer the richest transcriptomes but cover far fewer cells, BIDCell balances coverage with high expression-profile concordance, and SCS excels in morphology and spatial-diversity, while expression purity shows internal trade-offs and is not tied to other metrics.
Ligand–receptor analysis revealed STARmap PLUS and Stereo-seq alignments with expected cell populations, whereas MERFISH often misassigned interactions due to broader signal dispersion, illustrating how segmentation interacts with platform design.
A patch-based processing strategy handled large datasets and ensured fair comparisons, restricting analyses to high-quality segmented cells that met thresholds.
Researchers provide practical guidance: match segmentation tools to tissue architecture and platform, with image-based methods like SAM performing well on visually distinct cells, while transcriptomic-integrating methods are more computationally heavy, aided by a decision tree for goals like comprehensive maps, rare-cell recovery, or high-confidence profiles.
The study tests include SAM, SAM2, BidCell, SCS, Baysor, Cellpose, CelloType, Swin Transformer, Cascade Mask R-CNN, among others; two methods were excluded from quantitative comparisons due to grid-like artifacts.
Noted limitations include incomplete technology coverage, platform designs and tissue composition confounding algorithm vs. technology effects, limited gene-set analyses to one melanoma dataset, and reliance on serial sections rather than true re-runs; future work calls for replicate benchmarking and broader platform coverage.
Overall, segmentation quality fundamentally constrains downstream spatial biology, and the work provides a reproducible Nextflow pipeline and framework for evaluating future methods under standardized conditions.
A key takeaway is that method choice should be context-dependent, guided by the biological questions and the most relevant metrics rather than chasing a single global score.
The evaluation uses a comprehensive CellSPA-based metric suite covering cell detection, transcript assignment, transcriptomic richness, gene concordance, morphology, cell-type composition, marker purity, and spatial diversity, all normalized to a 0–1 scale.
When adjusting for coverage, SCS leads in balanced segmentation quality and cell yield, with BIDCell and Baysor close behind, and BIDCell and Baysor accumulating the most dataset wins.
Summary based on 2 sources
Get a daily email with more AI stories
Sources
Nature • Oct 23, 2026
Benchmarking segmentation methods for spatial transcriptomics
BIOENGINEER.ORG • Oct 23, 2026
Segmentation choices reshape spatial transcriptomics results, benchmark